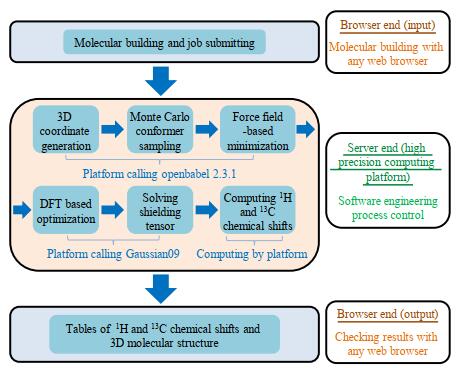
近年来,随着计算化学理论的不断突破,以及计算硬件性能的大幅提高,基于第一性原理针对有机分子开展1H和13C核磁共振化学位移的精确计算技术也取得了很大进展,部分方法已经逐步用于较大分子及较复杂分子体系的准确预测.本文建立了一个基于密度泛函理论的高精度有机分子化学位移在线计算系统平台,能够在线交互式完成分子量小于800的分子的化学位移计算服务.该系统平台工具能加快构建分子结构与核磁共振谱图的映射关系过程,为有机分子核磁共振谱图的高效指认,以及结构精确解析提供一个新的有力支撑.
李骞
,
唐亚林
,
向俊锋
. 基于密度泛函理论的高精度有机分子化学位移在线计算系统平台[J]. 波谱学杂志, 2021
, 38(1)
: 22
-31
.
DOI: 10.11938/cjmr20202831
With continuing breakthroughs in computational chemistry theory and substantial improvement of computation hardware performance, great progresses have been made in recent years in first principle-based prediction of 1H and 13C chemical shifts of organic molecules. Some methods have even been gradually applied for accurate prediction in complex molecular systems. In this paper, a density functional theory-based high-precision on-line chemical shift prediction platform for organic molecules is established, which provides on-line interactive service of chemical shift prediction for molecules with a molecular weight less than 800. The platform accelerates the mapping between nuclear magnetic resonance (NMR) spectra and molecular structures, and provides a powerful tool for efficient assignment of NMR spectra and accurate analysis of organic molecular structure.
[1] ABRAHAM R J, MOBLI M. Modelling 1H NMR spectra of organic compounds:theory, applications and NMR prediction software[M]. Hoboken, NJ:Wiley, 2008.
[2] BEEBY J, STERNHEL S, HOFFMANN T, et al. Estimation of chemical-shifts of aromatic protons using additive increments[J]. Anal Chem, 1973, 45(8):1571-1573.
[3] MATTER U E, PASCUAL C, PRETSCH E, et al. Estimation of chemical shifts of olefinic protons using additive increments.3. Examples of utility in nmr studies and identification of some structural features responsible for deviations from additivity[J]. Tetrahedron, 1969, 25(9):2023-2034.
[4] KUHN S, SCHLORER N E. Facilitating quality control for spectra assignments of small organic molecules:nmrshiftdb2-a free in-house NMR database with integrated LIMS for academic service laboratories[J]. Magn Reson Chem, 2015, 53(8):582-589.
[5] CASABIANCA L B, DE DIOS A C. Ab initio calculations of NMR chemical shifts[J]. J Chem Phys, 2008, 128(5):052201.
[6] BANFI D, PATINY L. www.nmrdb.org: Resurrecting and processing NMR spectra on-line[J]. Chimia, 2008, 62(4):280-281.
[7] MESTRENOVA, MESTRELAB RESEARCH S L[CP]. Santiago de Compostela, Spain.
[8] ChemOffice[CP]. CambridgeSoft Corp, Cambridge, MA, USA 02140-9802.
[9] ACD[CP]. Advanced Chemistry Development, Toronto, Ontario, Candada.
[10] BALLY T, RABLEN P R. Quantum-chemical simulation of 1H NMR spectra. 2. Comparison of DFT-based procedures for computing proton-proton coupling constants in organic molecules[J]. J Org Chem, 2011, 76(12):4818-4830.
[11] WILLOUGHBY P H, JANSMA M J, HOYE T R. A guide to small-molecule structure assignment through computation of (H-1 and C-13) NMR chemical shifts[J]. Nat Protoc, 2014, 9(3):643-660.
[12] SEFZIK T H, TURCO D, IULIUCCI R J, et al. Modeling NMR chemical shift:A survey of density functional theory approaches for calculating tensor properties[J]. J Phys Chem A, 2005, 109(6):1180-1187.
[13] LODEWYK M W, SIEBERT M R, TANTILLO D J. Computational prediction of H-1 and C-13 chemical shifts:A useful tool for natural product, mechanistic, and synthetic organic chemistry[J]. Chem Rev, 2012, 112(3):1839-1862.(标度参数详见http://cheshireNMR.info)
[14] HE X, WANG B, MERZ K M. Protein NMR chemical shift calculations based on the automated fragmentation QM/MM approach[J]. J Phys Chem B, 2009, 113(30):10380-10388.
[15] SWAILS J, ZHU T, HE X, et al. AFNMR:automated fragmentation quantum mechanical calculation of NMR chemical shifts for biomolecules[J]. J Biomol Nmr, 2015, 63(2):125-139.
[16] HE X, ZHU T, WANG X W, et al. Fragment quantum mechanical calculaton of proteins and its applications[J]. Accounts Chem Res, 2014, 47(9):2748-2757.
[17] ZHU T, ZHANG J Z H, HE X. Automated fragmentation QM/MM calculation of amide proton chemical shifts in proteins with explicit solvent model[J]. J Chem Theory Comput, 2013, 9(4):2104-2114.
[18] ZHU T, HE X, ZHANG J Z H. Fragment density functional theory calculation of NMR chemical shifts for proteins with implicit solvation[J]. Phys Chem Chem Phys, 2012, 14(21):7837-7845.
[19] HALGREN T A. MMFF VII. Characterization of MMFF94, MMFF94s, and other widely available force fields for conformational energies and for intermolecular-interaction energies and geometries[J]. J Comput Chem, 1999, 20(7):730-748.
[20] HALGREN T A. MMFF VI. MMFF94s option for energy minimization studies[J]. J Comput Chem, 1999, 20(7):720-729.
[21] PORCO J A, SU S, LEI X G, et al. Total synthesis and structure assignment of (+)-hexacyclinol[J]. Angew Chem Int Edit, 2006, 45(35):5790-5792.
[22] RYCHNOVSKY S D. Predicting NMR spectra by computational methods:Structure revision of hexacyclinol[J]. Org Lett, 2006, 8(13):2895-2898.
[23] Database for Organic Compounds[OL]. SDBS, https://sdbs.db.aist.go.jp.